Microbial growth depends heavily on physicochemical limits that dictate how cells consume surrounding nutrients. Over long evolutionary timescales, microorganisms have refined how they allocate protein resources to survive across diverse environmental conditions.
The study, titled Evolution Shapes Enzyme Turnover Numbers to Support Cellular Objectives, was published in the journal Nature.
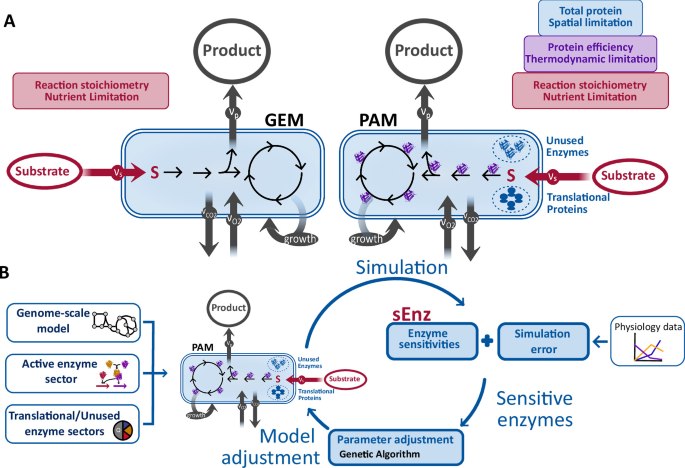
The newly introduced computational pipeline, known as the PAMparametrizer, splits its workflow into preprocessing, parametrization, and analysis steps. During preprocessing, researchers initialize coarse-grained protein sectors, including translational proteins and unused enzymes, using quantitative proteomics or data from related model organisms like E. coli.
After initial setup, the framework optimizes enzyme turnover numbers, or $k_{cat}$ values, to match experimentally observed metabolic fluxes. Parameter optimization proceeds through four distinct stages: selecting the most influential enzymes, optimizing their $k_{cat}$ values via a genetic algorithm, reparameterizing the model, and evaluating overall fitness.
To reduce computational complexity, the system selects enzymes based on sensitivity coefficients calculated using a tool called sEnz. These coefficients measure how shifts in enzyme concentration impact the overall cellular growth rate. A genetic algorithm then adjusts the turnover numbers to minimize the distance between model simulations and experimental data.
| Organism | Primary Focus | Observed Outcome |
|---|---|---|
| Escherichia coli | Metabolic behavior | Captured traits improved |
| Corynebacterium glutamicum | Protein allocation | Captured traits improved |
| Pseudomonas putida | Extracellular phenotype | Limited data recovered phenotype |
When applied to organisms like Escherichia coli and Corynebacterium glutamicum, the optimized Protein Allocation Models captured metabolic behavior more accurately than initial configurations. For the metabolically versatile bacterium Pseudomonas putida, limited experimental data restricted results to the recovery of the extracellular phenotype.
The research team combined the flux control quantification method with genetic algorithms to link exchange fluxes directly to enzyme kinetics. This approach aims to establish a scalable foundation for evolution-informed model parametrization across various organisms and environmental conditions.
Related reading