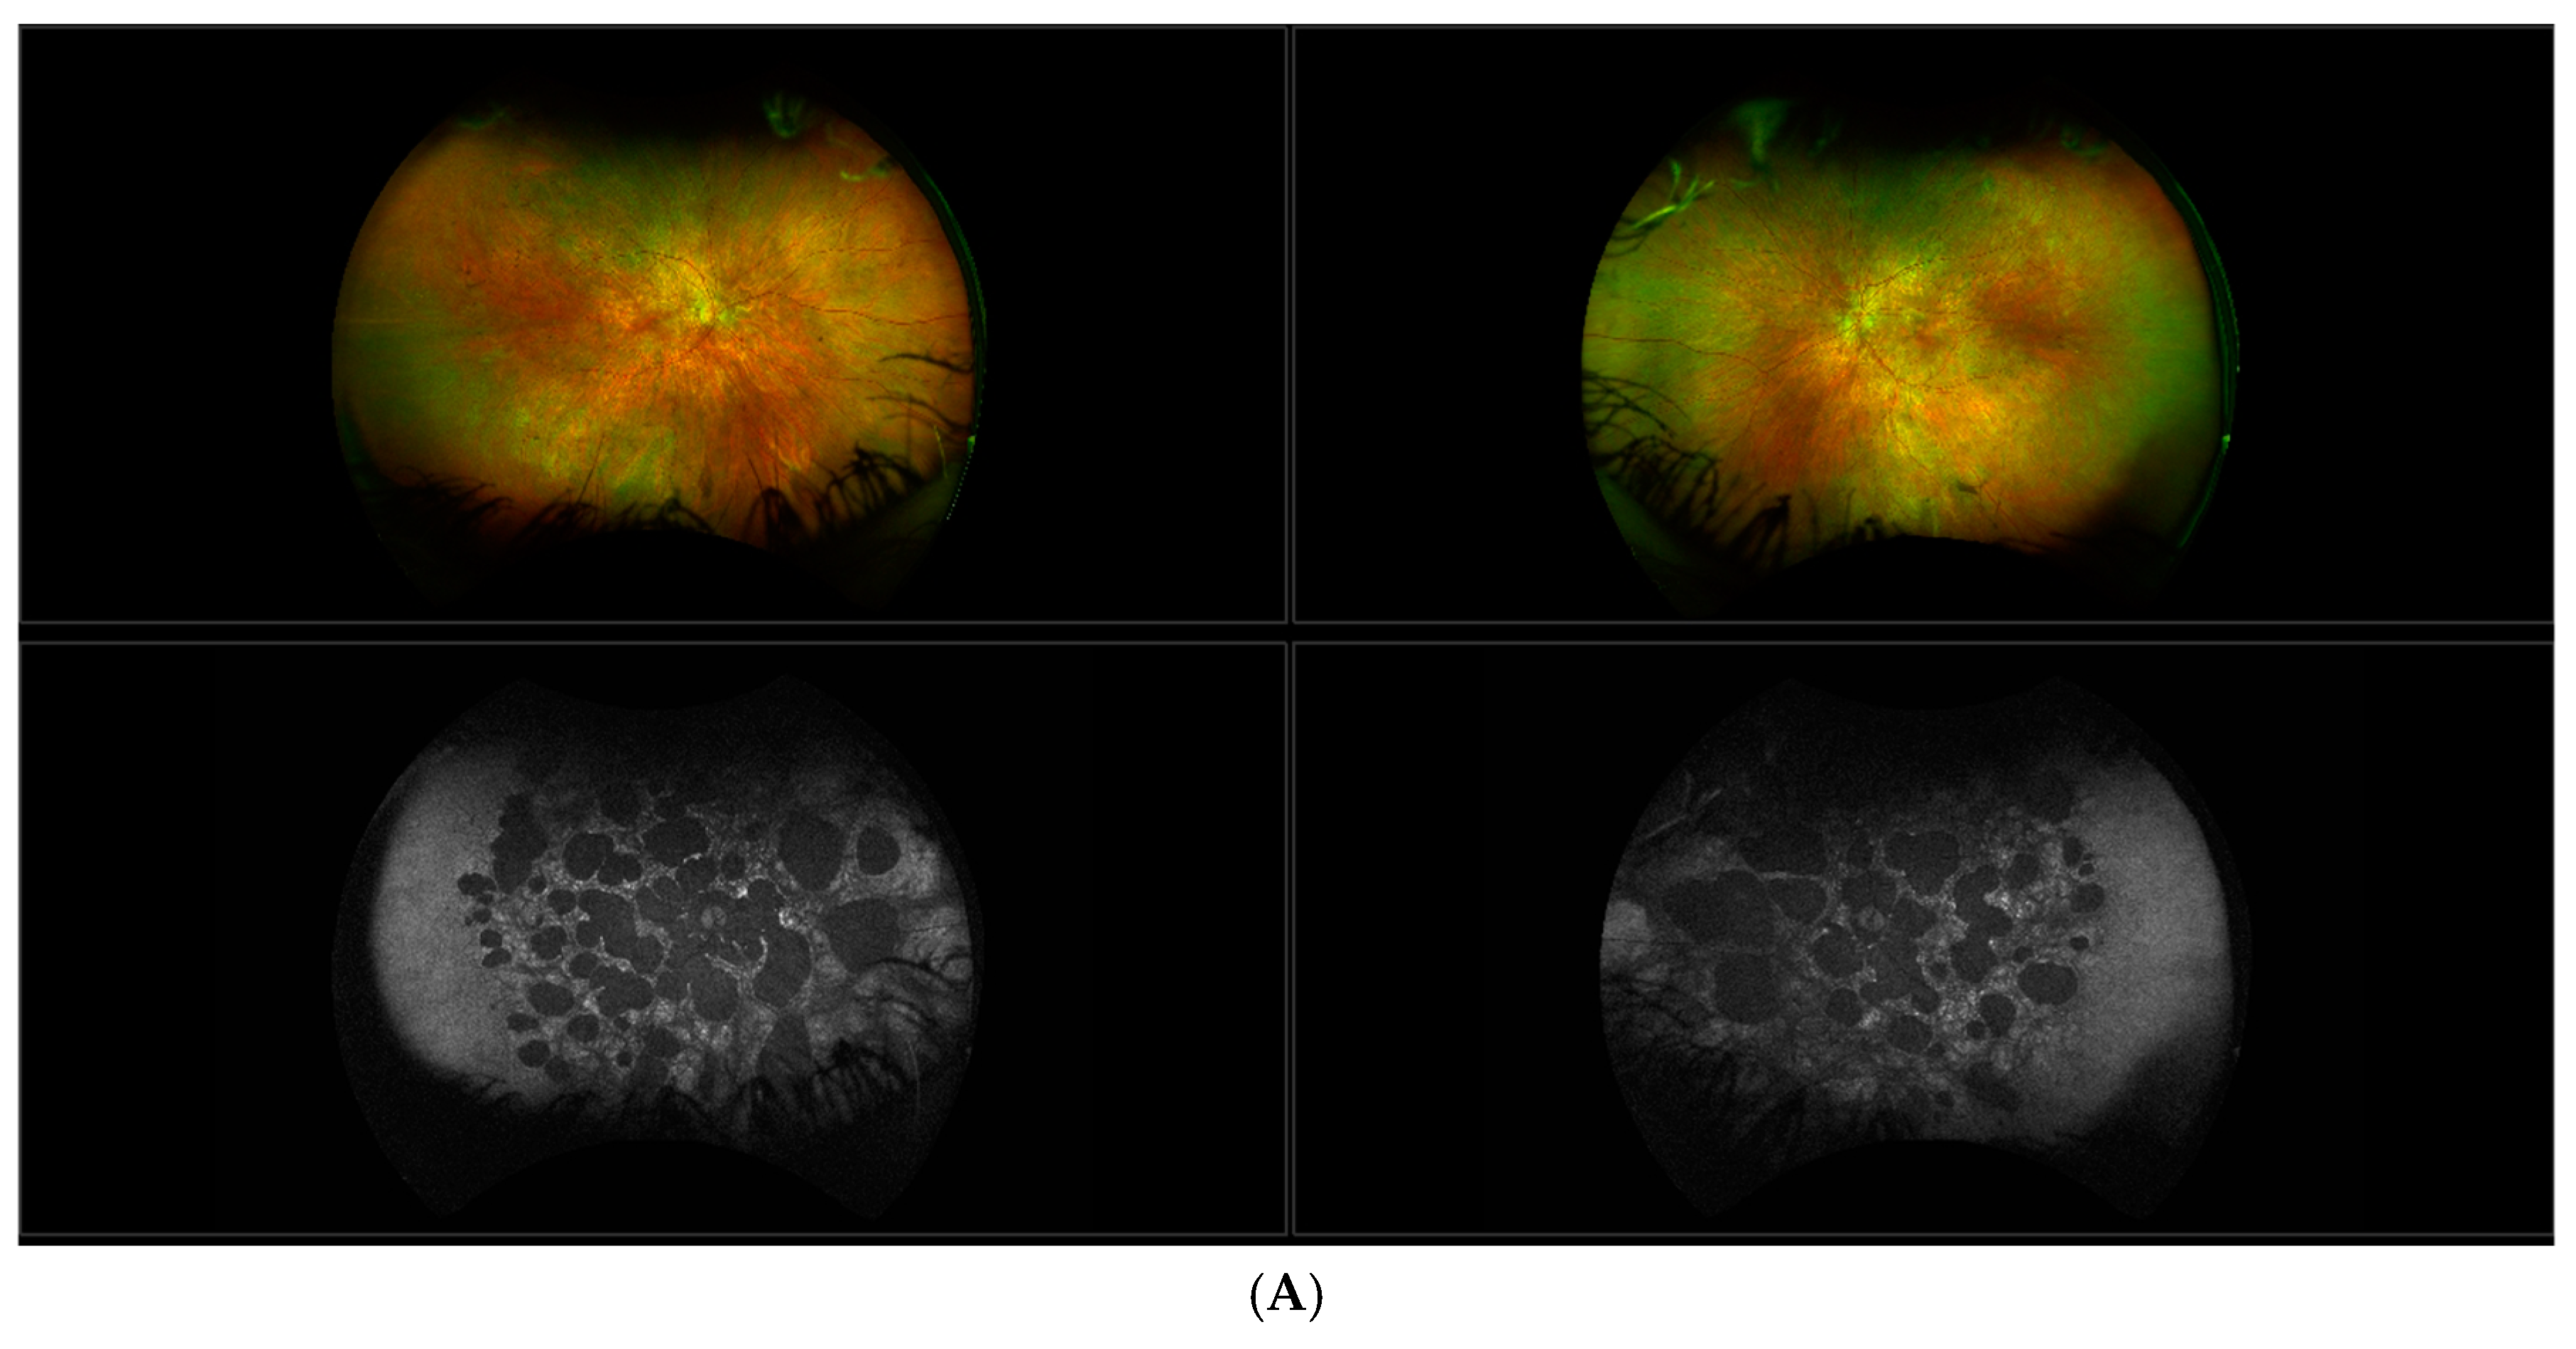
Retinitis pigmentosa caused by biallelic variants in the RPE65 gene—known as RPE65-RP—leads to severe visual impairment by early adulthood, according to a retrospective observational study of nine patients across four Japanese institutions.
Clinical Progression and Visual Impairment
Patients diagnosed with RPE65-RP typically exhibit symptoms beginning in early childhood, generally between the ages of 2 and 7, though adult-onset cases also occur. Clinical assessments conducted across the study group, including best-corrected visual acuity (BCVA) and electroretinogram testing, revealed that while five patients initially maintained visual acuity of 20/40 or better, the condition is progressive. Seven of the nine patients eventually reached a BCVA of 20/250 or worse in at least one eye. The study noted that none of the participants exhibited nystagmus, a rhythmic, involuntary eye movement often associated with early-onset retinal conditions.
Variability in Retinal Degeneration
The study highlighted that the severity of vision loss can vary based on specific genetic variants. Two patients who carried compound heterozygous variants—specifically c.1543C > T (p.Arg515Trp) and c.683A > C (p.Gln228Pro)—demonstrated a slower rate of progression. These individuals retained better visual acuity and showed wider residual ellipsoid zones and visual fields compared to other participants. Despite these differences, all nine patients displayed retinal degeneration consistent with retinitis pigmentosa, characterized by non-detectable responses during electroretinogram testing.
Study Methodology and Scope
Researchers evaluated clinical data from seven unrelated individuals and one pair of siblings to characterize the phenotypic features of the disease. The diagnostic process involved a combination of fundus and autofluorescence imaging, optical coherence tomography, and Goldmann perimetry to map the extent of the retinal damage. The findings confirm that RPE65-RP serves as a significant cause of visual handicap, with the majority of patients facing profound vision loss by the time they reach early adulthood.
Frequently Asked Questions
What is the primary cause of vision loss in RPE65-RP?
The condition is caused by biallelic variants in the RPE65 gene, which is essential for the visual cycle in the retina. Mutations in this gene prevent the proper regeneration of visual pigments, leading to the progressive degeneration of photoreceptor cells.
How does the age of onset affect the prognosis?
While most patients experience symptoms in early childhood, the rate of degeneration varies. Some patients with specific genetic variants may maintain relatively good vision for a longer period, but the study indicates that most individuals will experience severe visual impairment by early adulthood.
Are there universal clinical signs for this condition?
All patients in the study showed retinal degeneration consistent with retinitis pigmentosa and non-detectable electroretinogram responses. However, clinical features such as nystagmus were absent in this cohort, demonstrating that the presentation can differ from other forms of inherited retinal dystrophy.